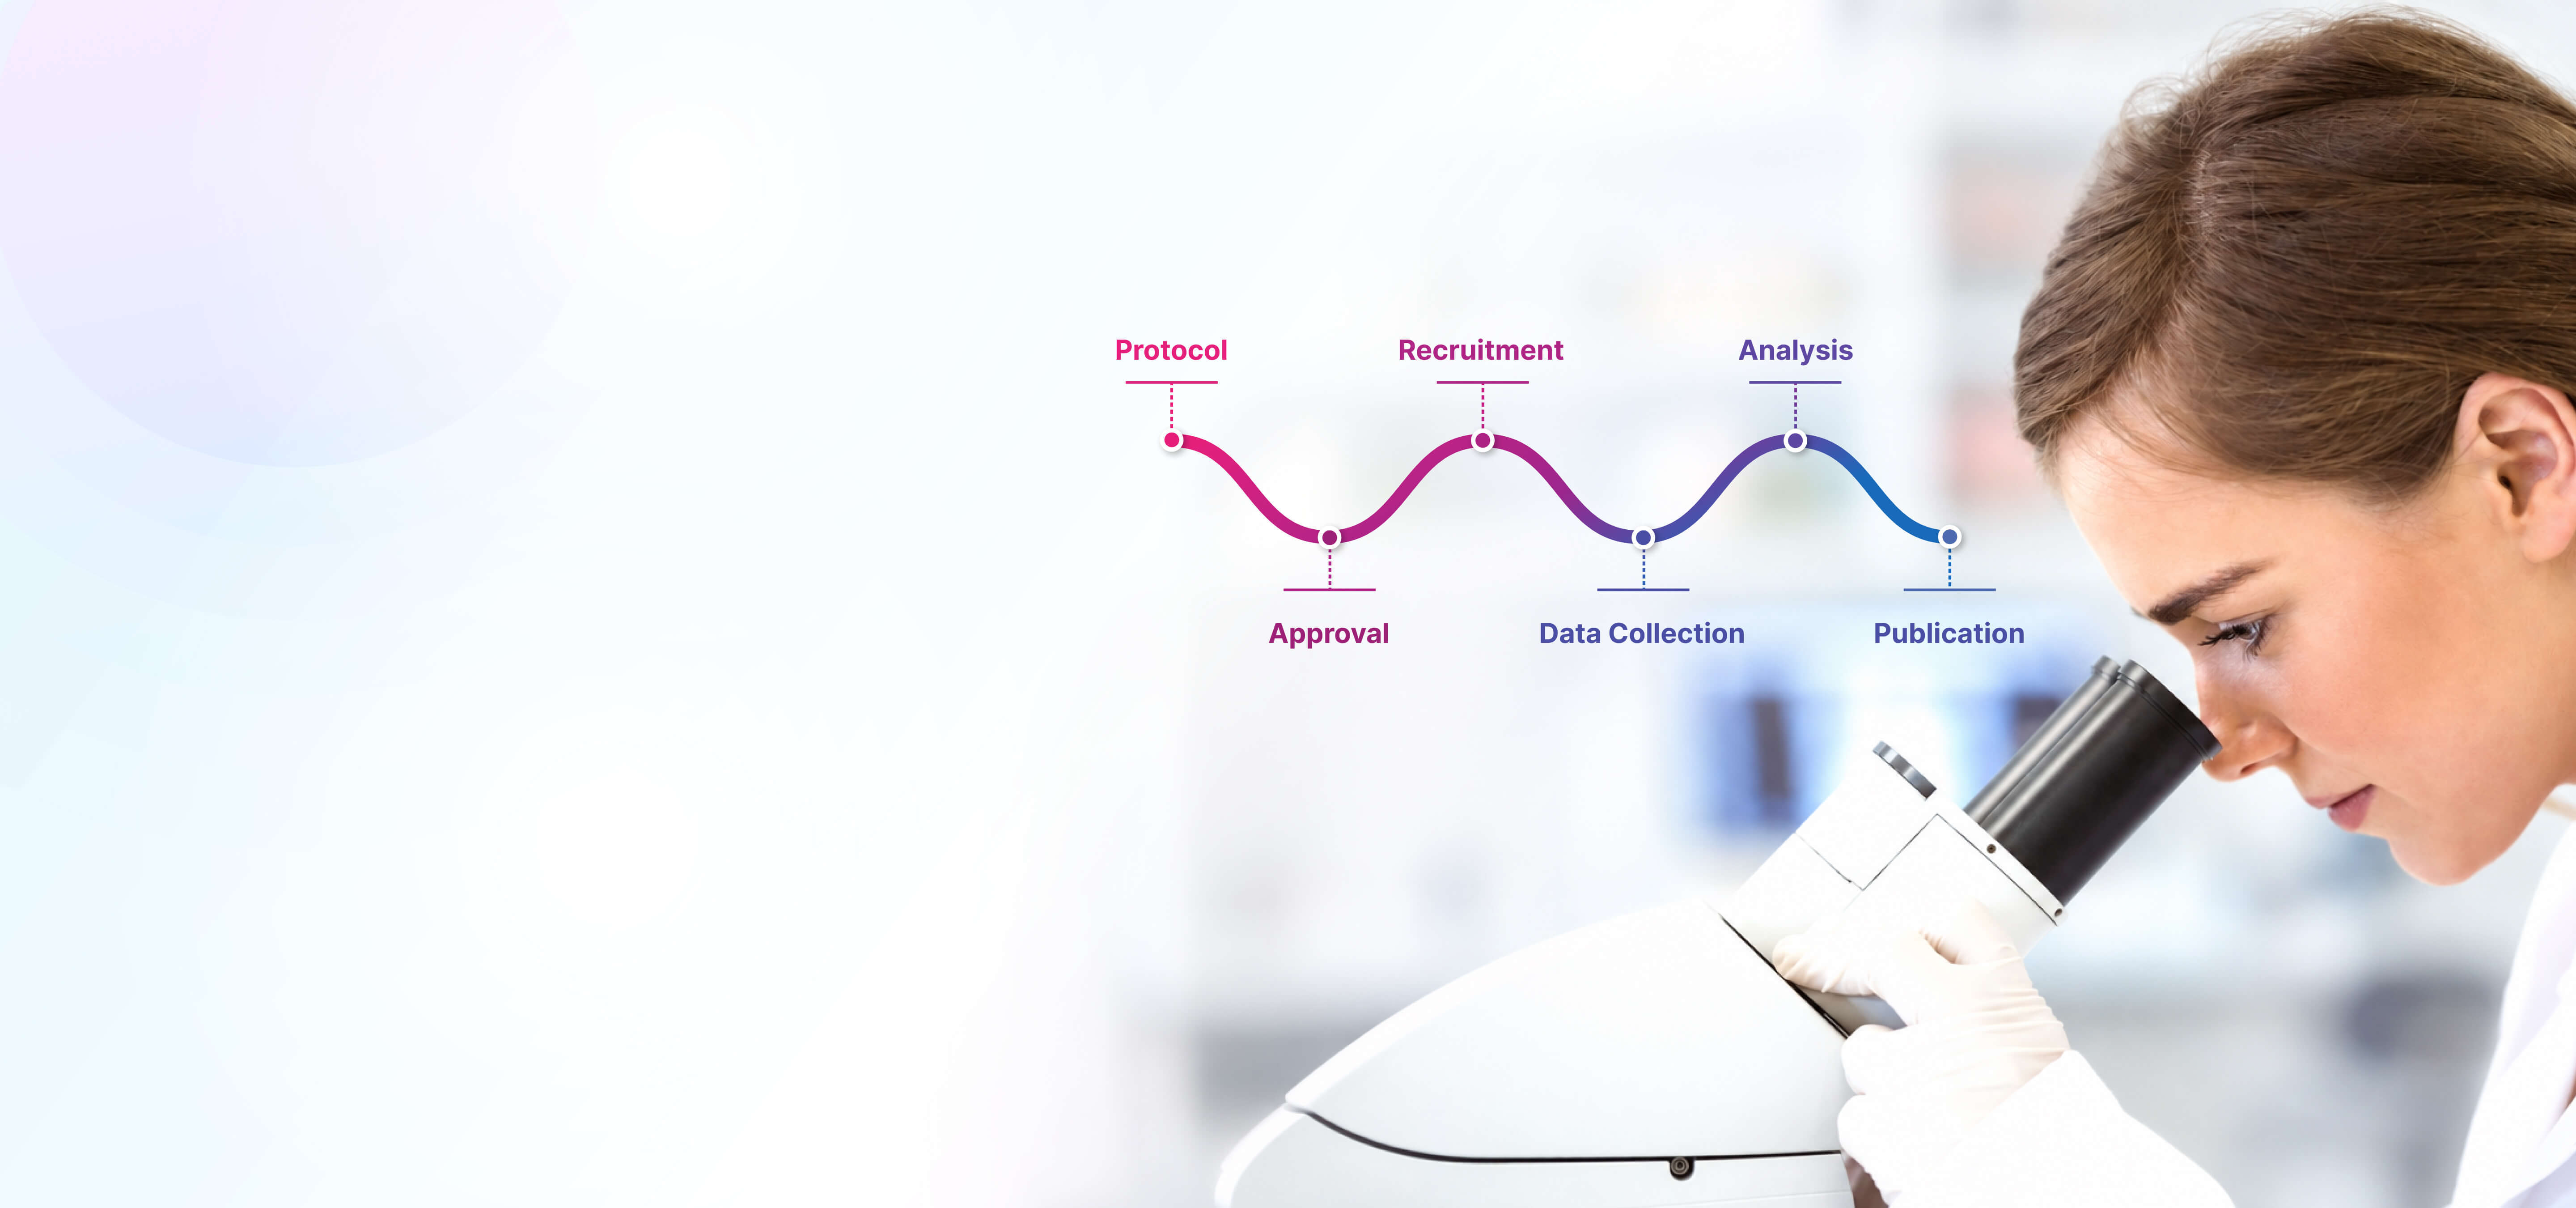
From protocol to publication
Regulators accept the conclusions they can trace. Alethium's statisticians and medical writers work directly from the study record, with every value traceable to the event that produced it, from the analysis plan through the clinical study report.

Regulatory Strategy
Submission planning across FDA, EMA, ANVISA, and IRB/EC pathways, matched to each market you file in.

IRB & Ethics Submissions
Initial submissions, amendments, and continuing review across central IRBs and ethics committees worldwide, prepared, tracked, and archived in one place.

Essential Documents & TMF
The trial master file maintained continuously and audit-ready, with every document versioned and traceable to the event that produced it.

Safety Reporting
SAE intake, expedited reporting to authorities and ethics committees, and safety-letter distribution, tracked to acknowledgment.
Consent Compliance
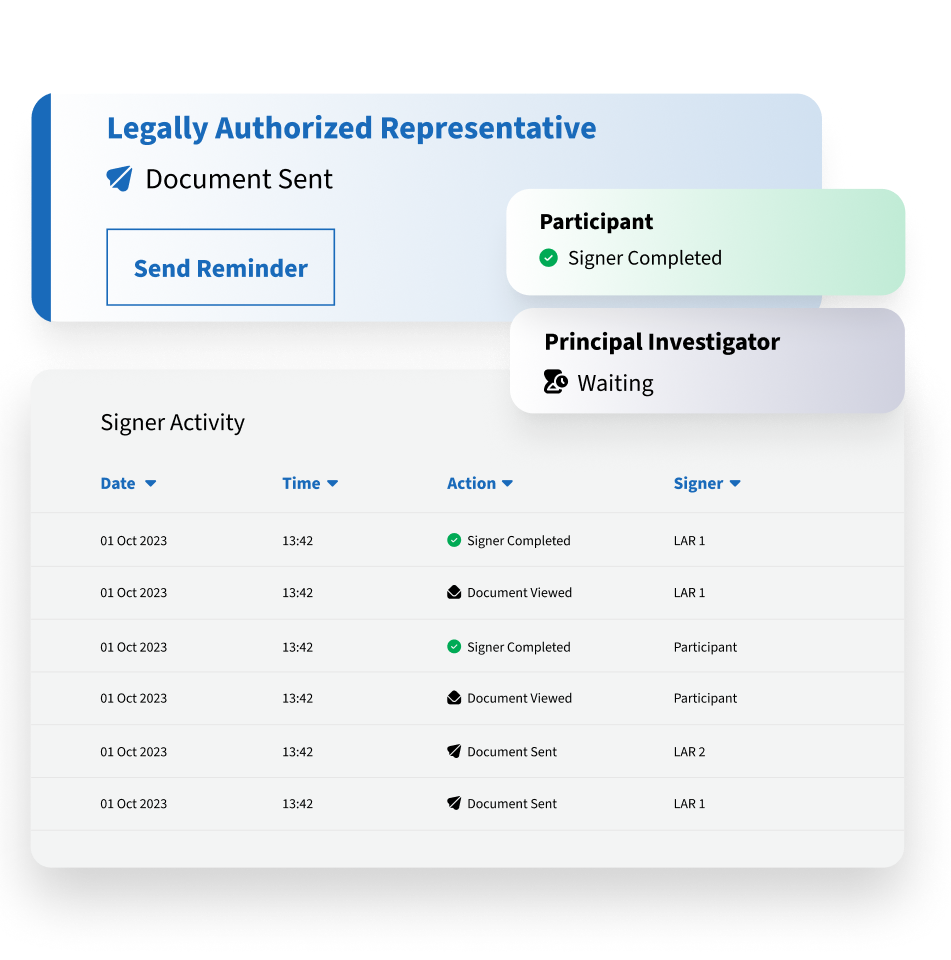
Part 11-compliant consent and automatic re-consent when protocols change.
Licensing & Import/Export
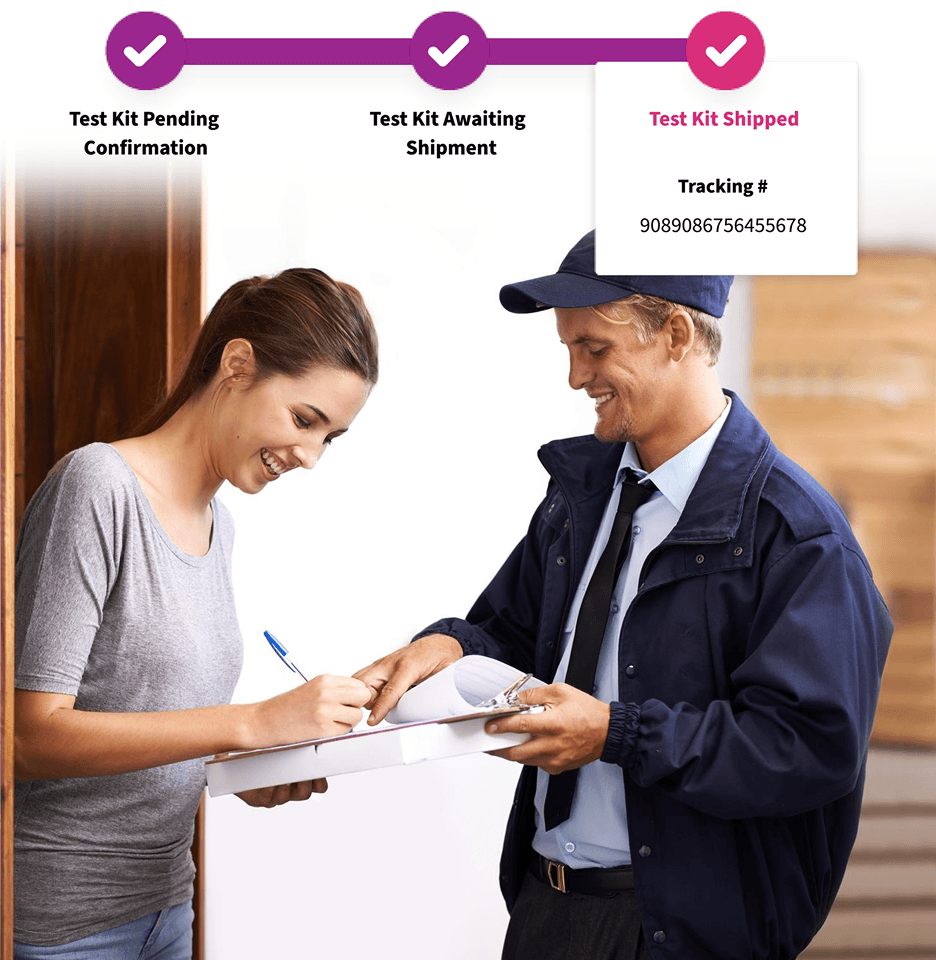
Import and export licensing, customs, and Importer of Record services live with the supply chain.
Audit Readiness
Once the study is running, evidence accumulates on its own. On Alethium's architecture the audit trail is not a report you assemble at closeout. It is the system of record. Every consent, shipment, and data point is already evidence, and the automated test suite doubles as a living traceability matrix.
Submitted. Approved. Audit-Ready.
Your regulatory department, from day one.
FAQs - Common Questions
Alethium manages the full regulatory pathway for a trial: submission strategy across FDA, EMA, ANVISA, and IRB/EC pathways; IRB and ethics submissions including amendments and continuing review; trial master file maintenance; safety reporting; and import, export, and Importer of Record services for study supplies.
The TMF is maintained continuously rather than assembled at closeout. Because every service runs on one event-driven platform, each document is versioned and traceable to the event that produced it — the consent, the shipment, the visit — so the evidence a regulator asks for already exists in the system of record.
Yes — that is the model Alethium is built for. Emerging biopharma companies face the same regulatory obligations as large sponsors with a fraction of the headcount. Alethium supplies the strategy, submissions, documentation, and safety reporting that an internal regulatory department would otherwise own.
Yes. Alethium prepares and tracks submissions to central IRBs and ethics committees across the markets a study runs in, including amendments, continuing reviews, and renewals. Region-specific requirements such as language translations and data residency assurances are managed as part of the submission package.
Regulatory responsibility for expedited safety reporting sits with the sponsor. Alethium operates the process on the sponsor's behalf: SAE intake, expedited reporting to authorities and ethics committees, and safety-letter distribution to sites, each tracked through to acknowledgment.
Part 11 requirements map directly to platform mechanisms: an immutable, time-stamped audit log; electronic signatures; document versioning; and role-based access controls. Protocol rules are expressed as Behavior-Driven Development scenarios validated by automated tests, so the test suite functions as a living traceability matrix.
